The future of sterile QA in biotech manufacturing will depend on how well quality teams evaluate changing processes and the evidence behind them. Technologies that reduce exposure or provide earlier information can help manufacturers, but each brings questions about suitability, control and human judgement. Our view is that QA needs to influence these decisions during process design, when changes remain easier to make.
Biotech manufacturing also covers different product types. Sterile biological medicines and advanced therapy medicinal products should not be treated as interchangeable. The European Commission lists Annex 2 for biological products and separate Part IV GMP guidance for advanced therapies. Understanding the applicable framework is an essential first step before borrowing practices from another manufacturing platform.
Barrier technologies change the questions QA must ask
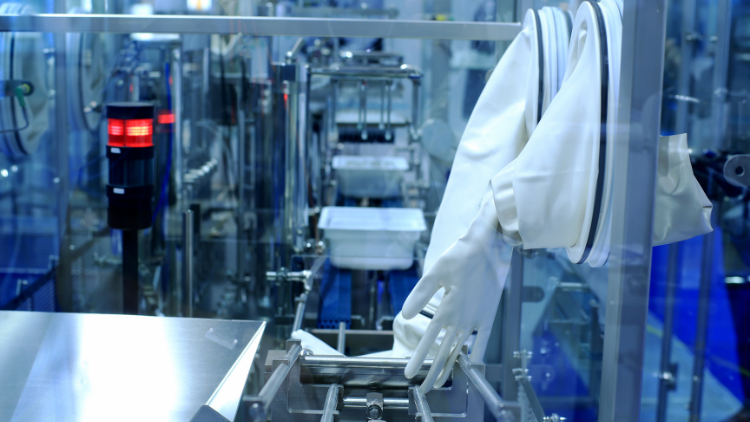
Annex 1 addresses isolators, restricted access barrier systems and the use of closed systems to protect critical operations. These approaches offer a basis for reducing exposure, but a technology’s name does not explain how protection is maintained during a particular manufacturing process.
As a practical recommendation, QA should assess the complete operational sequence: preparation, transfer, processing, intervention and recovery from an interruption. A process may be well protected during routine production yet depend on a more exposed activity during troubleshooting. That activity needs explicit consideration when the equipment and procedures are being selected.
This has implications for investment decisions. When reviewing a proposed platform, ask the project team to demonstrate how a representative batch and a credible interruption would be managed. Involving operators and maintenance specialists makes the discussion more useful than relying solely on the supplier’s description of normal operation.
Faster microbiology needs a defined purpose
FDA’s production and process control questions and answers encourage beneficial microbiological testing innovations, including rapid methods. The agency also distinguishes the limitations of aseptic process monitoring from the direct evidence available for terminal sterilisation. Faster information does not provide a general permission to dispense with required sterility assurance controls.
For QA teams, a method’s proposed use should determine the evaluation. An early process warning, an organism identification and a result supporting product release answer different questions. Before implementation, agree what decision the result will support, how uncertainty will be handled and how the method’s performance will be demonstrated for that use.
A useful adoption project begins with a clearly described problem. If the objective is earlier investigation, measure whether the new information changes the response time and quality of the investigation. If it supports release, involve regulatory colleagues early to understand the authorisation and submission implications. The technology should earn its place through relevant evidence.
Advanced therapies require product-specific judgement
Some advanced therapies have limited sample availability or short shelf lives that complicate conventional testing timelines. EU Part IV guidance addresses adapted sterility assurance strategies and validated rapid microbiological methods where justified. It also requires appropriate mitigation when sterility results are unavailable at release. These provisions belong to the specific ATMP framework and authorisation context.
The practical challenge is to define the decision process before a time-sensitive batch reaches the point of release. Teams should know which information must be available, who evaluates it and how they respond to a result received later. Rehearsing a hypothetical adverse result can reveal unclear responsibilities or communication gaps without waiting for a live event.
This also changes how QA capacity is planned. A small batch may need intensive coordination because its patient, testing and delivery arrangements are specific. Resourcing decisions should therefore consider the complexity of each quality decision alongside production volume.
Digital oversight must preserve accountable decisions
Connected manufacturing systems can make it easier to compare processing events with quality observations. As an editorial recommendation, manufacturers should begin with a defined use case, such as improving review of recurring interruptions, and establish whether the necessary data is sufficiently complete and consistent.
The Commission’s proposed Annex 22, published for consultation in 2025, addresses AI model selection, validation, monitoring and human oversight. The current EudraLex listing does not present it as an operative annex. This distinction matters when discussing the future: proposed guidance signals regulatory priorities, while current obligations and the intended use of a system determine today’s controls.
For an AI-assisted quality tool, a practical evaluation should ask what happens when the tool misses an important event or produces an incorrect recommendation. Human reviewers need enough context to challenge its output. A confident-looking summary is of limited value if the team cannot check the underlying evidence.
Prepare QA expertise alongside manufacturing capability
Sterile QA in biotech manufacturing will require teams that can connect microbiological understanding with validation, process knowledge and data interpretation. Training should use realistic decisions, including ambiguous results and interrupted operations, so that staff practise applying judgement under the conditions they may encounter.
QA Resources supports biologics manufacturers through quality assurance, validation, regulatory and quality systems services. Contact the team to discuss specialist support for a new platform, a technology transfer or a review of the quality capability required as your sterile manufacturing processes develop.